Back to:
| Ordered alphabetically by REFCODE | Ordered alphabetically by Common name | Ordered by molecular formula | Grouped by family |
Olanzapine
(Last updated October 2024)
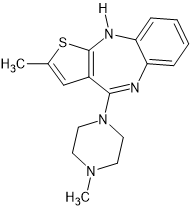
Figure 1. The molecular diagram of Olanzapine.
CSP studies
This work was carried out in collaboration with Susan Reutzel-Edens and Rajni Miglani-Bhardwaj when they were working at Eli Lilly and the University of Strathclyde, respectively. A rigid molecule MOLPAK search was carried out in 2010, but this has not been published. This is called search B. A flexible CrystalPredictor search was carried out in 2013. This is called search A.
There were second derivative properties which could not be uploaded to the database, but the free energy corrections derived from these could.
| REFCODE |
| ||
| Formula | C17 H20 N4 S1 | ||
| Common Name | Olanzapine | ||
| IUPAC Systematic Name | 2-Methyl-10-(4-methyl-1-piperazinyl)-4H-3-thia-4,9-diaza-benzo[f]azulene | ||
| Other Names | |||
| CSD Refcodes | UNOGIN03, UNOGIN04, UNOGIN06, UNOGIN05 | ||
| Search Identifier |
| ||
| Scientist | Louise Price | ||
| Date | 2013 | ||
| Publication | Bhardwaj RM, Price LS, Price SL, Reutzel-Edens SM, Miller GJ, Oswald IDH, Johnston B, Florence AJ 2013. Cryst Growth Des 13, 1602-1617. | ||
| Energy model |
| ||
| Study_ID | 30 | ||
| Programs | Study_ID=11, DMACRYS (2.0.4) | ||
| Location on S Drive | \CHEMISTRY_CPOSS\Olanzapine\PCM | ||
| Potential Description | GDMA2.2(PCMdielectric3(PBE0/6-31G(d,p))) + FIT with isotropic S | ||
| Intramolecular Description | PBE0/6-31G(d,p) | ||
| Energy model |
| ||
| Study_ID | 11 | ||
| Programs | Study_ID=20, CrystalOptimizer, DMACRYS (2.0.4) | ||
| Location on S Drive | \CHEMISTRY_CPOSS\Olanzapine\CrystalOptimizer | ||
| Potential Description | GDMA2.2(PBE0/6-31G(d,p)) + FIT with isotropic S | ||
| Intramolecular Description | PBE0/6-31G(d,p) | ||
| Energy model |
| ||
| Study_ID | 20 | ||
| Programs | Flexible CrystalPredictor (1.x), dmaflex-Quick, DMACRYS (2.0.4) | ||
| Location on S Drive | \CHEMISTRY_CPOSS\Olanzapine\CrystalPredictor | ||
| Potential Description | GDMA2.2(MP2/6-31G(d,p))multipoles rotated from gas phase local minimum + FIT with isotropic S | ||
| Intramolecular Description | SCF/6-31G(d,p) | ||
| Search Identifier |
| ||
| Scientist | Louise Price | ||
| Date | 2010 | ||
| Publication | Early search - not published | ||
| Energy model |
| ||
| Study_ID | 10 | ||
| Programs | Study_ID=0, dmaflex, DMAREL (4.1.1) | ||
| Location on S Drive | \CHEMISTRY_CPOSS\0-EarlySearches\home\louise_price.eminerals\Olanzapine_dmaflex | ||
| Potential Description | SCF 6-31G(d,p DMA(v1.2)+ fit.pots+iso S | ||
| Intramolecular Description | SCF | ||
| Energy model |
| ||
| Study_ID | 0 | ||
| Programs | Molpak, DMAREL (4.1.1) | ||
| Location on S Drive | \CHEMISTRY_CPOSS\0-EarlySearches\home\louise_price.eminerals\olanzapine | ||
| Potential Description | SCF 6-31G(d,p) DMA(v1.2)+ fit.pots+iso S | ||
| Intramolecular Description | MP2/6-31G(d,p) | ||
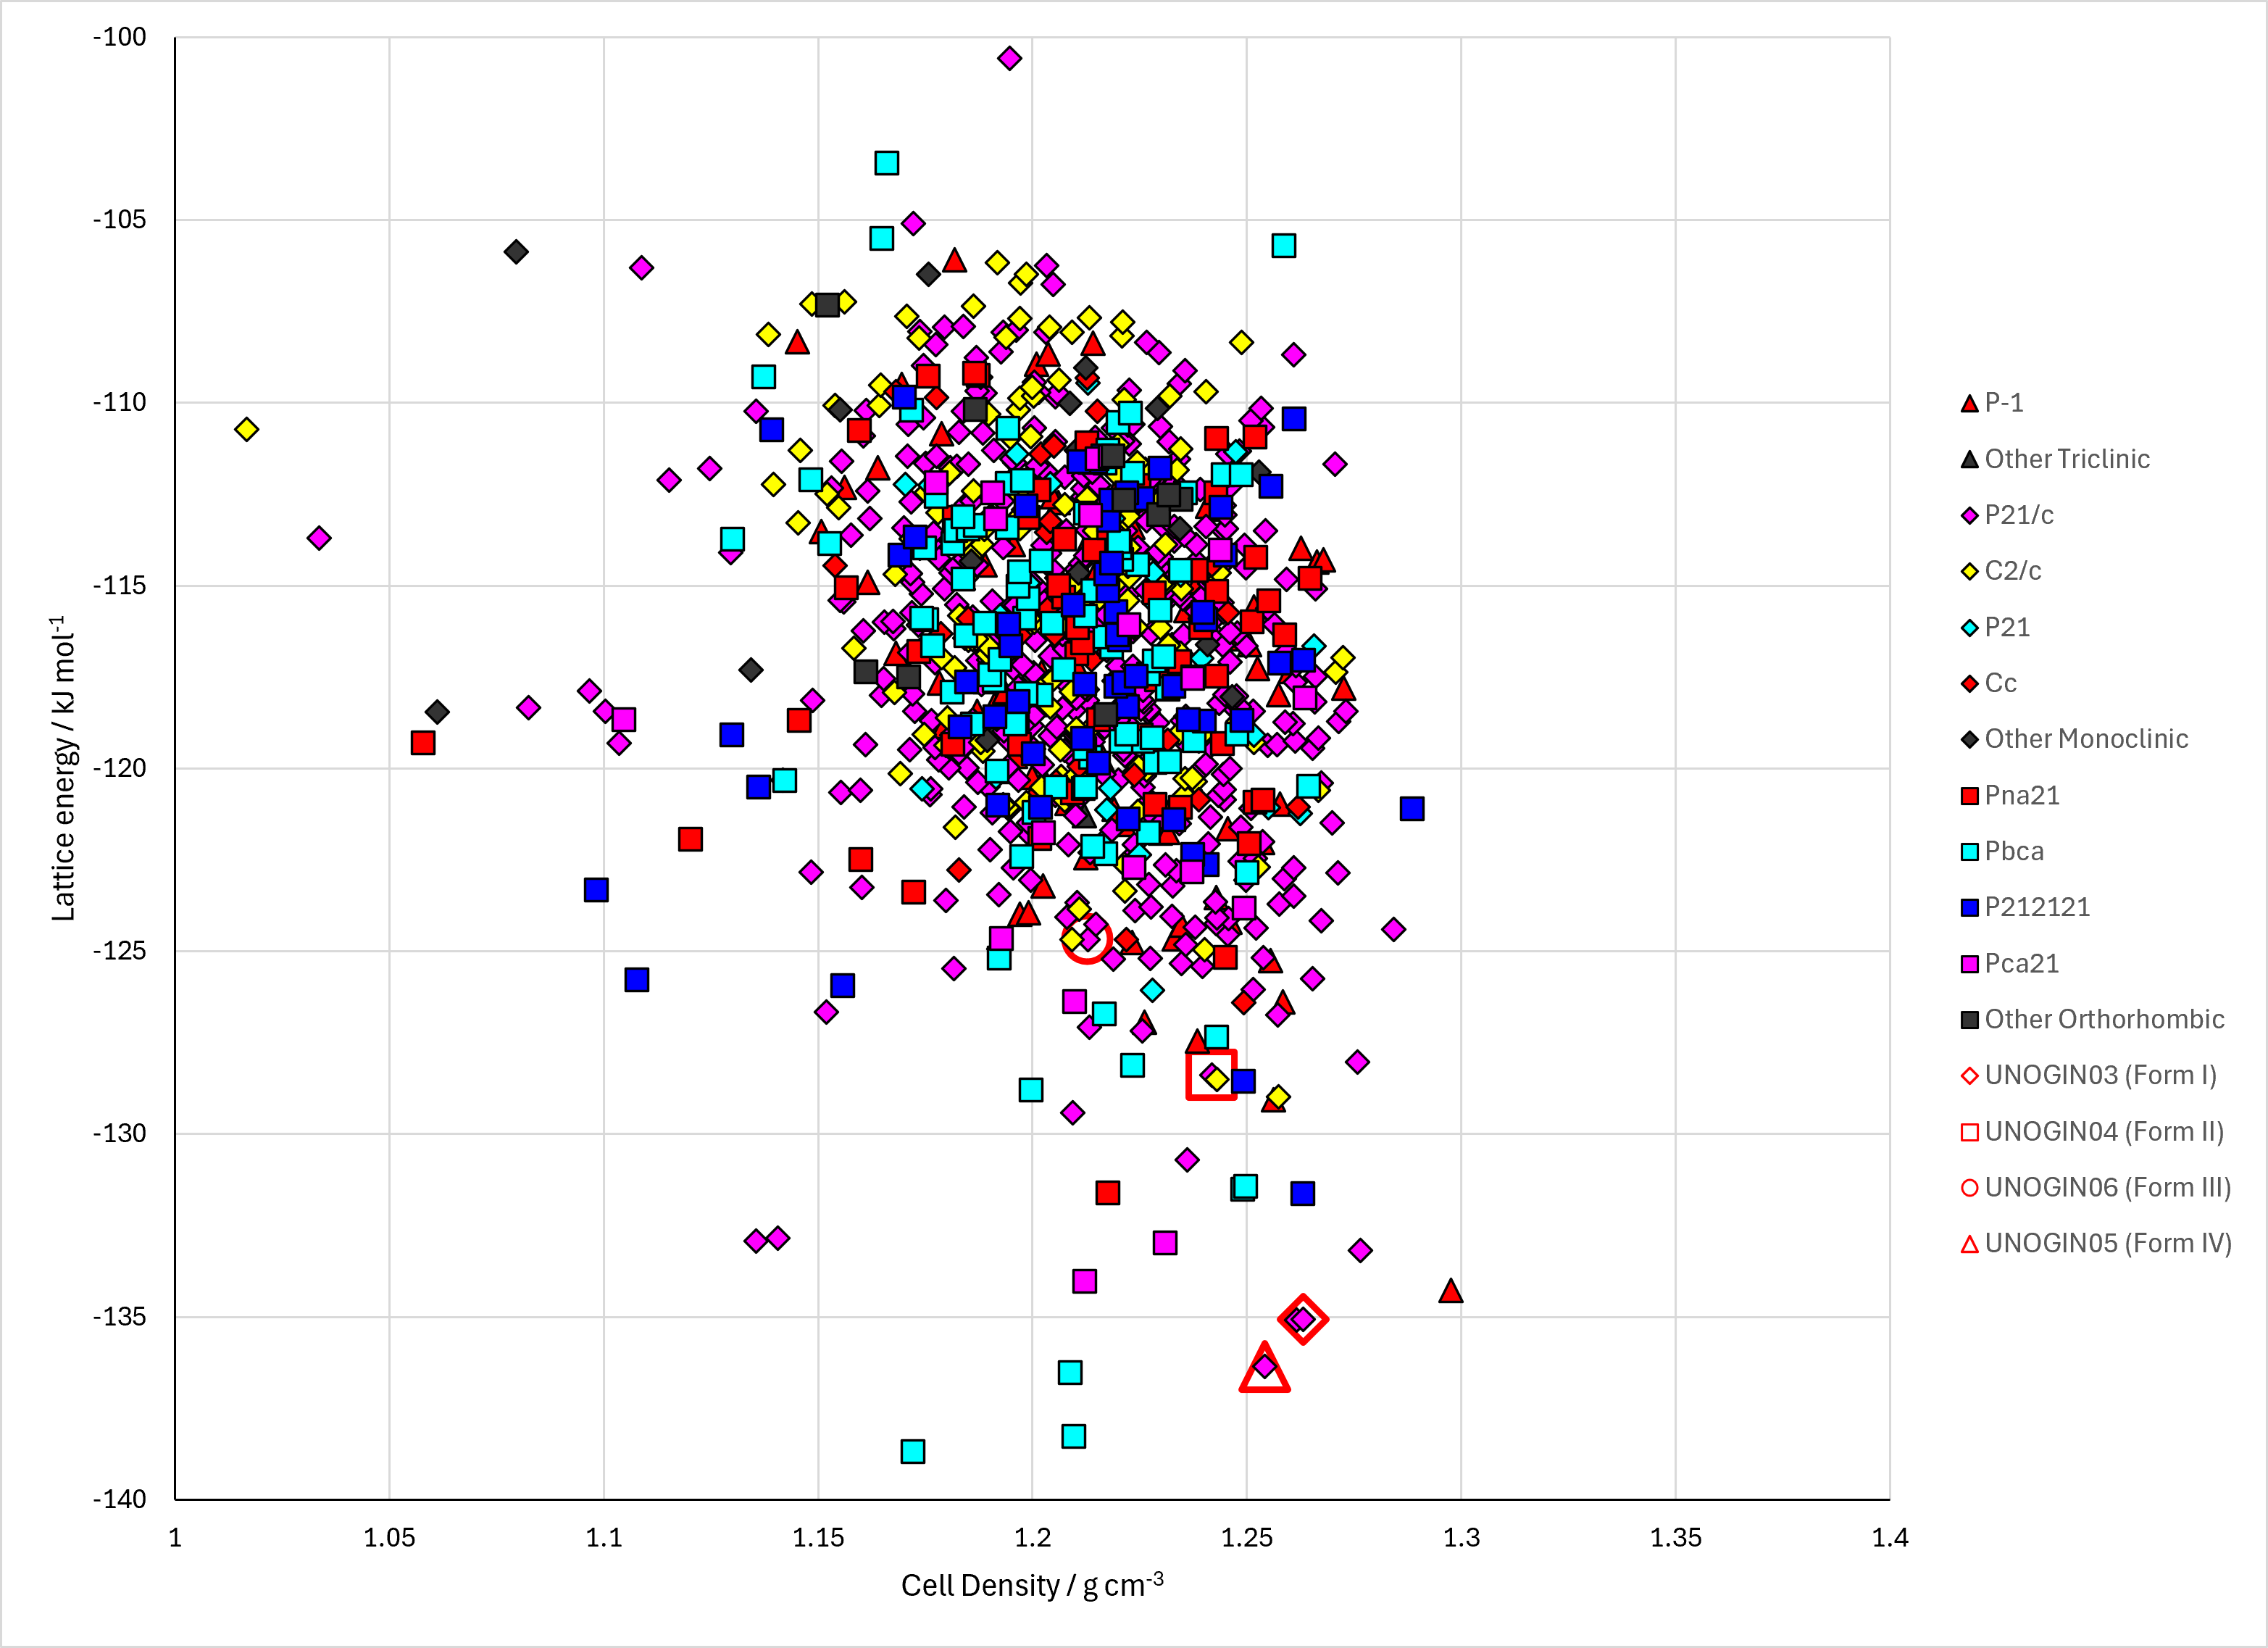
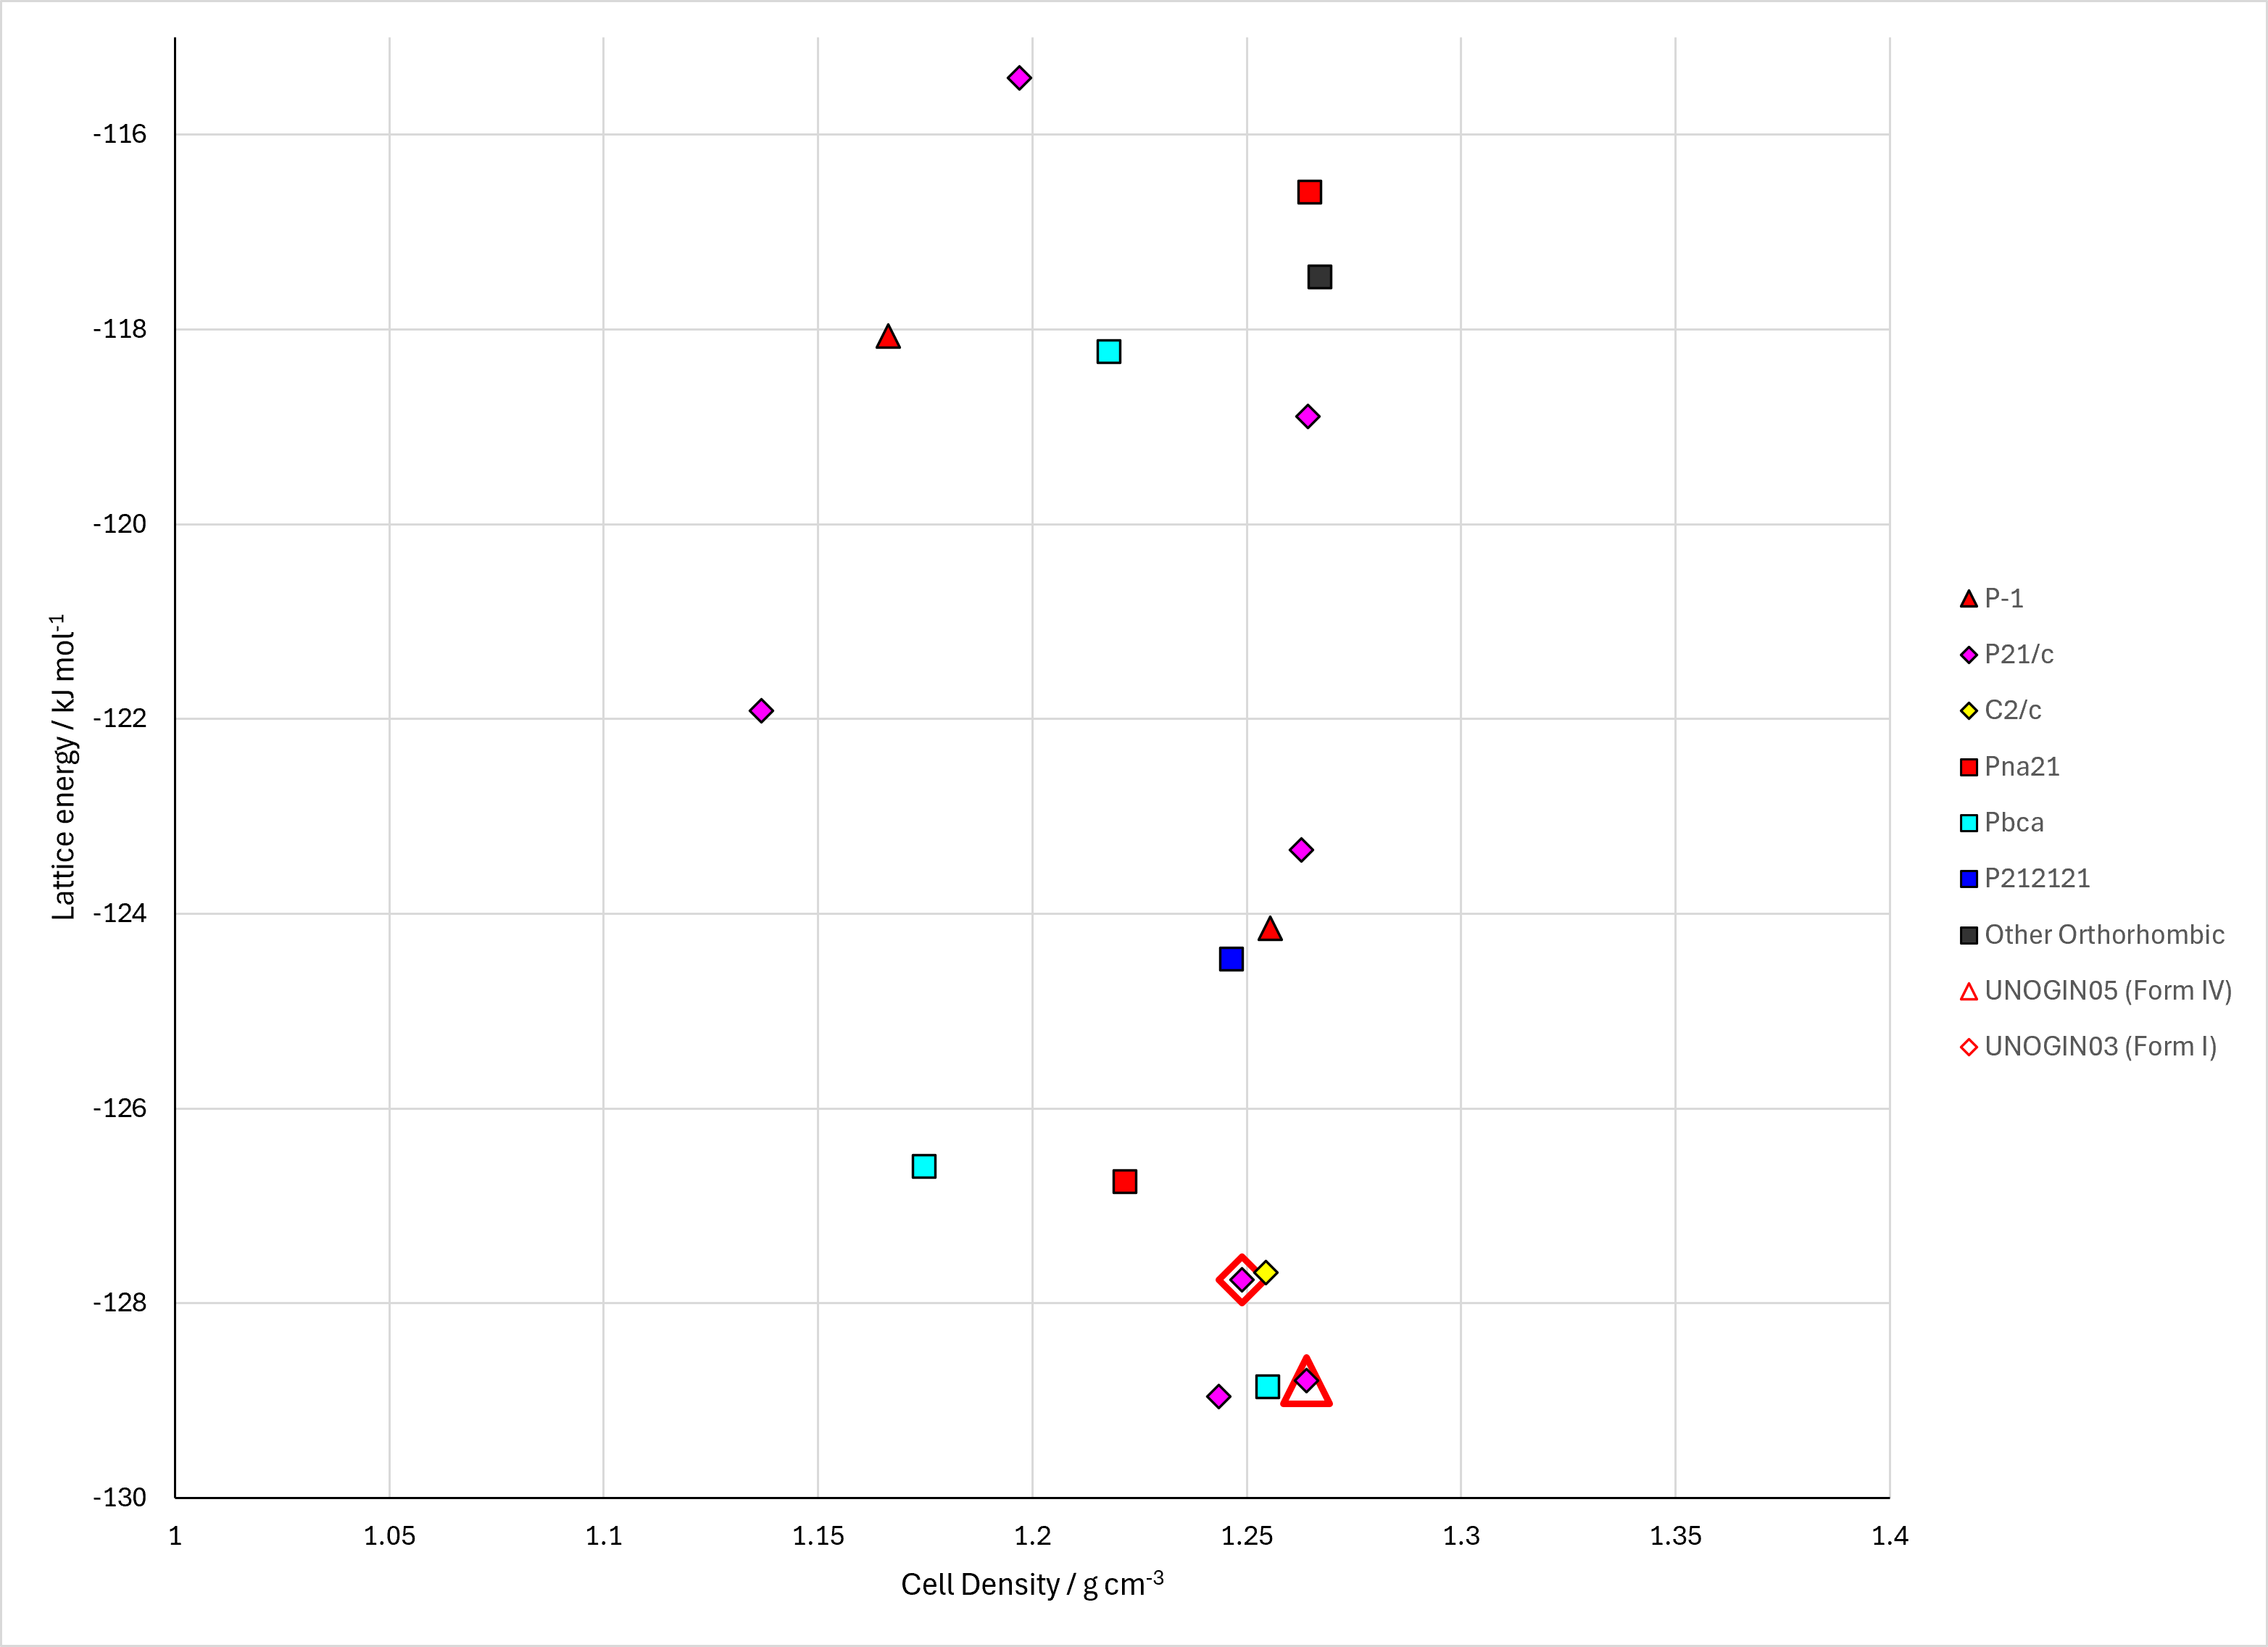
Figure 2. Lattice Energy Landscapes of Olanzapine. Energy models A3 and B2.
CSD structures (CSD version 5.44 with Jun and Sep 2023 updates)
Table 1. Crystallographic information and experimental information of CSD entries for Olanzapine. Different polymorphs, as ascertained by structure matching of 20 molecule coordination sphere in Mercury, are coloured differently. Reference polymorphs are outlined in bold.
| REFCODE | Spacegroup | Z' | a / Å | b / Å | c / Å | α / ° | β / ° | γ / ° | density / g cm-3 | Form | R factor | T / K | Year | Comment / reference | REFCODE |
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| UNOGIN | P21/c | 1.0 | 10.388 | 14.839 | 10.567 | 90.0 | 100.64 | 90.0 | 1.2963 | stable polymorph I | 3.33 | at 293 K | 2004 | 10.1107/S160053680302806X | UNOGIN |
| UNOGIN01 | P21/c | 1.0 | 10.3830 | 14.826 | 10.56 | 90.0 | 100.616 | 90.0 | 1.2988 | stable polymorph I | 4.3 | at 293 K | 2003 | 10.1021/cg034055z | UNOGIN01 |
| UNOGIN02 | P21/c | 1.0 | 9.913 | 16.5329 | 9.9992 | 90.0 | 98.023000 | 90.0 | 1.2788 | polymorph II | 4.66 | at 298 K | 2011 | 10.1107/S0108270111043952 | UNOGIN02 |
| UNOGIN03 | P21/c | 1.0 | 10.3411 | 14.521 | 10.5314 | 90.0 | 100.291 | 90.0 | 1.3336 | polymorph I | 3.34 | 123 K | 2013 | 10.1021/cg301826s | UNOGIN03 |
| UNOGIN04 | P21/c | 1.0 | 9.8544 | 16.314 | 9.9754 | 90.0 | 98.304 | 90.0 | 1.3077 | polymorph II | 5.42 | 123 K | 2013 | 10.1021/cg301826s | UNOGIN04 |
| UNOGIN05 | P21/n | 1.0 | 8.6555 | 15.4441 | 12.5558 | 90.0 | 95.284 | 90.0 | 1.2416 | polymorph IV | 10.5 | 443 K | 2019 | 10.1021/acs.cgd.8b01881 | UNOGIN05 |
| UNOGIN06 | P21/c | 1.0 | 10.708 | 16.476 | 10.065 | 90.0 | 110.43 | 90.0 | 1.2471 | polymorph III | 11.39 | 293 K | 2024 | 10.1107/S2052252524007383 | UNOGIN06 |
Crystallization conditions are included in the downloadable Word Document.
Other Notes
The study was originally done with two conformational regions named eq(uatorial) and ax(ial) relating to the manner in which the benzodiazepine group linked to the nitrogen atom. Susan Reutzel-Edens was not happy with these terms being used for a nitrogen atom, so they were changed to A and B respectively. (A is the conformation seen in the experimental crystal structures and is the lower in energy.) Crystal structures have been given to other groups with the old names, and hence these are occasionally referred to in the literature.
A162 was originally thought to be the best match for form III, although Rajni Miglani Bhardwaj noted that there were problems with it and it was not perfect which is why it is referred to it as a model for form III in that paper.
© UCL Chemistry Department 2022. This page was last updated on 30 Sep 2026. If you have any problems with this page please email the WebMaster